Dwara
Justin Wong
—
Regulatory Medical Writing: Key Documents aur Unhe Kaise Likhein

Justin Wong
Vikas Prabhari
Graduated kiya Bachelor's me Global Business & Digital Arts, Minor me Entrepreneurship
Regulatory medical writing ko lagbhag har jagah ek career ke roop mein bataya jata hai aur shayad hi kahi deliverable documents ke ek set ke roop mein. Aap is role ke baare mein teen lambe guides padh sakte hain aur fir bhi aapko nahi pata chalega ki aapko kaun se documents produce karne honge, ya kaun sa guideline aapko batata hai ki har ek ko kaise structure karna hai.
Yeh us problem ko solve karta hai. Neeche har ek document ka map diya gaya hai, jisme governing guideline aur har item ke liye ek link hai, saath hi submission structure kaise kaam karta hai, 2025 mein sach mein kya badla hai, aur review cycle asal mein kaise chalta hai.
<CTA title="Draft Technical Documents With Fewer Loose Ends" description="Jenni helps you structure long documents and keep terminology consistent across every section" buttonLabel="Try Jenni Free" link="https://app.jenni.ai/register" />
Regulatory Medical Writing Kya Hai
Regulatory medical writing un documents ka production hai jo medicines ya device regulator ko bheje jaate hain: jaise FDA, EMA, MHRA, PMDA aur unke counterparts. Yeh kisi product ke life cycle ke shuruat se lekar aakhir tak chalta hai, yaani pehla protocol likhne se lekar (kisi ko dose dene se pehle) marketing application tak, aur un periodic safety reports tak jo tab tak file ki jaati hain jab tak product market mein hai.
Is kaam ko do features define karte hain. Pehla, structure aapke shuru karne se pehle hi tay hota hai, ek aisi guideline dwara jo public hai aur padhne ke liye free hai. Aur doosra, reader ek regulatory reviewer hota hai jiska professional farz hai inconsistency ko pakadna. Yeh combination is kaam ko expression se zyada accuracy, traceability aur yeh janne ke baare mein banata hai ki aapke saamne jo document hai us par kaun si rulebook apply hoti hai.
<ProTip title="🗺️ Orientation:" description="The fastest way to look competent in a regulatory writing interview is to name the guideline that governs each document. That knowledge is free and most candidates have not bothered" />
CTD, aur Har Document ka Ek Module Number Kyun Hota Hai
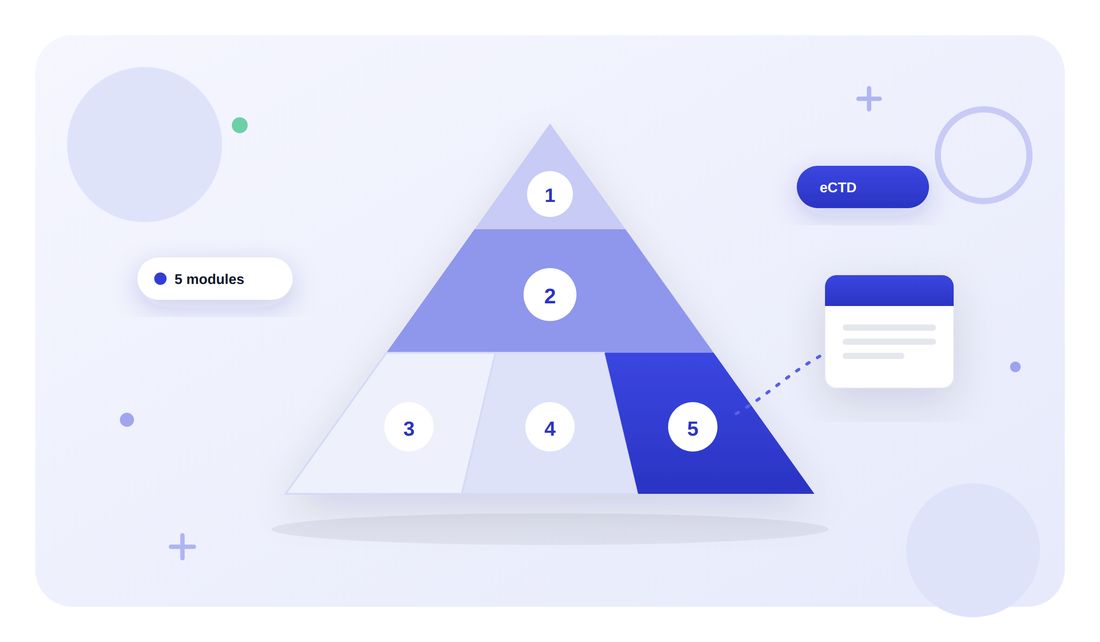
Marketing applications ko ek Common Technical Document (CTD) ke roop mein organize kiya jata hai, jise ICH M4 dwara define kiya gaya hai. Aap jo kuch bhi likhte hain woh iske paanch modules mein kahi na kahi aata hai, aur regulatory baatein document ke naam ke bajaye module numbers mein ki jaati hain.
Module 1, administrative information aur prescribing information. Region specific hota hai, aur technically yeh CTD ka hissa nahi hai.
Module 2, summaries. Yeh woh jagah hai jahan sabse senior regulatory writing hoti hai. Isme Quality Overall Summary, Nonclinical Overview aur clinical writers ke liye sabse zaroori, Clinical Overview (2.5 par) aur Clinical Summary (2.7 par) shaamil hote hain.
Module 3, quality. Chemistry, manufacturing, aur controls.
Module 4, nonclinical study reports.
Module 5, clinical study reports. Individual trial reports yahan rehte hain.
Practice mein sabse mahatvapurna antar Module 5 aur Module 2 ke beech hai. Ek clinical study report ek trial ko describe karti hai. Clinical Overview (2.5) sabhi trials ke aadhar par product ke baare mein argument karta hai, aur Clinical Summary (2.7) sabooto ko ikatha karta hai, jisme 2.7.3 efficacy ko cover karta hai, 2.7.4 safety ko, aur 2.7.6 individual studies ke synopses ko collect karta hai. Unki content requirements ICH M4E se aati hain, na ki ICH E3 se.
Is sab ke liye electronic format eCTD hai. FDA ise CDER aur CBER ke submissions ke liye standard format ke roop mein describe karta hai, aur yeh new drug applications, abbreviated applications, biologics license applications, aur commercial investigational new drug applications ke liye zaroori hai. September 2024 se Version 4.0 ko support kiya gaya hai.
Woh Documents Jo Aap Sach Mein Likhenge
Yeh rasta (map) hai. Inme se har ek guideline padhne ke liye free hai, aur aapke agle document ko govern karne wali guideline ko padhna aapke liye sabse kimti samay hoga.
Document | Yeh kya hai | Kiske dwara govern hota hai |
Clinical trial protocol | Ek trial ka plan: design, population, treatments, endpoints, statistics | ICH E6(R3), Annex 1, Appendix B |
Investigator brochure | Product ke baare mein sab kuch jo pata hai, jo trial chalane wale logon ke liye compile kiya gaya hai | ICH E6(R3), Annex 1, Appendix A |
Informed consent form | Ek participant ko trial mein hissa lene ke liye haan kehne se pehle kya bataya jata hai | ICH E6(R3) section 2.8, aur US mein 21 CFR 50.25 |
Clinical study report | Ek complete trial ka pura hisab, solah sections mein | |
Clinical Overview (2.5) aur Clinical Summary (2.7) | Ek marketing application mein product level argument aur evidence assembly | |
Plain language summary of results | Trial ke results ka ek aam bhasha (lay) summary, jo EU mein zaroori hai | EU Regulation 536/2014 Article 37, Good Lay Summary Practice ke saath |
Periodic benefit risk evaluation report | Ek marketed product ke liye chal rahi safety reporting | |
Development safety update report | Abhi bhi development mein chal rahe product ke liye annual safety reporting | |
Clinical evaluation report | Europe mein ek medical device ke liye clinical evidence dossier | EU MDR 2017/745, Article 61 aur Annex XIV, working template ke roop mein MEDDEV 2.7/1 revision 4 ke saath |
Briefing documents aur meeting packages | Kisi agency meeting ya advisory committee ke liye taiyaar ki gayi samagri | Koi ek single guideline nahi hai. Procedure specific, jo agency aur meeting type dwara tay hoti hai |
Inme se teen par dhyan dena zaroori hai. Plain language summary is field ka sabse underrated document hai: EU ko trial khatam hone ke barah mahine ke andar ek summary chahiye hoti hai, pediatric trials ke liye cheh mahine, aur Good Lay Summary Practice guidance lagbhag 12 saal ya usse bade readers ke liye readability target set karti hai. Isko achhi tarah likhna sach mein mushkil hai aur yeh skill har jagah kaam aati hai.
Informed consent form woh jagah hai jahan regulatory writing real logon se milti hai. E6(R3) simple aur concise language ki maang karta hai, aur 21 CFR 50.25 aath basic elements ke saath cheh additional elements set karta hai. Yeh chota hota hai, iski gahri janch hoti hai, aur yeh shuruat ke liye ek achha assignment hai.
Briefing documents is rule ke exception hain: inko koi ICH guideline govern nahi karti, isliye inka structure agency, procedure aur purani traditions se aata hai. Pediatric investigation plans aur orphan designation applications ke liye bhi yahi sach hai, jo ICH ke bajaye regional regulations dwara govern hote hain.
<ProTip title="🔗 Traceability:" description="Every number you write should be traceable to a source document in one step. If you cannot say which table or report a figure came from, you do not yet know whether it is right" />
Regulatory Writing Medical Communications Se Kaise Alag Hai
Log aksar in dono ke beech bina yeh samjhe choose karte hain ki dono ke din kitne alag hote hain.
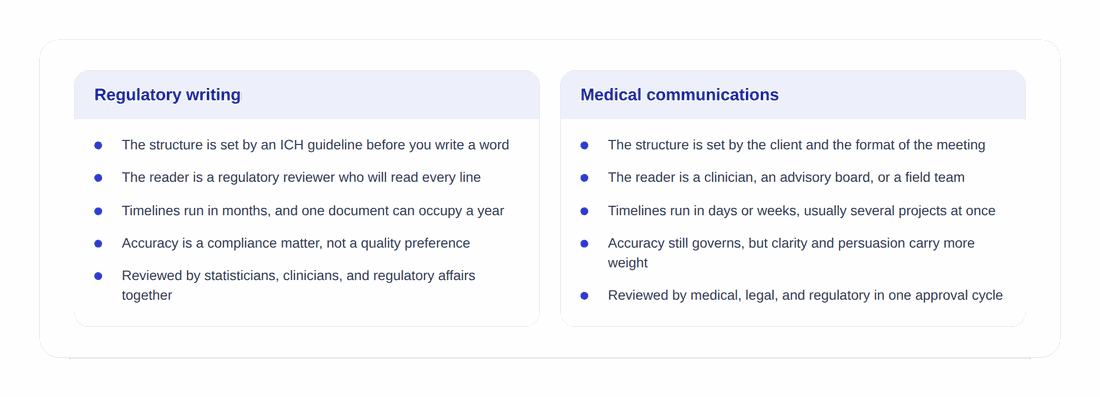
Regulatory writing mein, aapke ek shabd likhne se pehle hi structure ek guideline dwara set hota hai, reader ek reviewer hota hai jo har ek line padhega, timelines mahino mein chalti hain aur ek single document ek saal le sakta hai, accuracy ek compliance ka mamla hai na ki quality preference ka, aur review statisticians, clinicians aur regulatory affairs ke saath ek hi kamre mein hota hai.
Medical communications mein, structure client aur meeting ke format dwara set hota hai, reader ek clinician ya advisory board ya field team hoti hai, timelines dino ya hafto mein chalti hain jisme ek sath kai projects chal rahe hote hain, clarity aur persuasion ko accuracy ke sath zyada mahtva diya jata hai, aur review ek single medical, legal aur regulatory approval cycle ke throw chalta hai.
Dono mein se koi bhi mushkil nahi hai. Yeh alag-alag temperaments ke hisab se suit karte hain. Regulatory sabr, precision aur procedure ke tolerance ko reward karta hai. Medical communications speed, range aur ambiguity ke sath comfort ko reward karta hai.
2025 Mein Kya Badla
Is subject par adhiktar published guidance iske pehle ki hai, jise janana zaroori hai agar aap purani material padh rahe hain.
ICH E6(R3) 6 January 2025 ko step 4 par pahunch gaya. Yeh Good Clinical Practice ka current version hai aur yeh guideline ko kaafi restructure karta hai, protocol aur investigator brochure content requirements ko Annex 1 appendices mein move karta hai. Koi bhi course ya article jo E6(R2) ko current batata hai, ab purana ho chuka hai.
SPIRIT 2025 April 2025 mein publish hua tha, jo ek sath Nature Medicine, The Lancet, JAMA, aur BMJ mein dikha. Yeh trial protocols ke liye reporting checklist ko update karta hai aur yeh woh reference point hai jise ab adhiktar journals aur sponsors regulatory requirement ke saath use karte hain.
eCTD v4.0 ko September 2024 se FDA dwara support kiya gaya hai. Yeh writing change ke bajaye ek submission format change hai, lekin yeh is baat ko affect karta hai ki documents ko kaise assemble aur version kiya jata hai, jo ki practice mein writer ke liye ek badi problem hai.
Review Cycle Asal Mein Kaise Kaam Karta Hai

Ek regulatory document likh kar turant approve nahi ho jata. Yeh ek loop se gujarta hai, aam taur par kai baar, aur isi loop mein naye writers apna samay kho dete hain.
Ek typical cycle aise chalta hai: team ke saath agree kiya gaya ek shell ya outline, ek defined review group ko bheja gaya first draft, ek fixed window ke andar mile comments, ek comment resolution meeting jahan contradictory comments ko ek kamre mein settle kiya jata hai, ek revised draft, source documents ke khilaf ek quality control check, aur aakhir mein sign off. Bade documents ke liye is sequence ka beech ka hissa do ya teen baar repeat hota hai.
Do habits un logon ko alag karti hain jo ise achhe se handle karte hain. Pehli, kabhi bhi aise comment ko accept na karein jise aap samajhte nahi hain. Agar koi clinician aise badlav ke liye kehta hai jo kisi statistician ke pehle comment ke khilaf hai, to jawab ek baat-cheet hai, na ki koi compromise phrasing jo dono mein se kisi ko satisfy na kare. Doosri habit hai version discipline. Yeh janna ki comment kis draft ke khilaf kiya gaya tha, aur aisa keh pane ki capacity, ek lambe review cycle mein sabse aam failure ko rokti hai, jo ki do versions baad kisi correction ka chupke se undo ho jana hai.
<ProTip title="📑 Version control:" description="Keep a short decision log alongside the document recording what was changed, why, and who asked. It takes minutes per round and it answers the question that always comes up three months later" />
Regulatory Writing Mein Kaise Kadam Rakhein
Sabse reliable entry point ek contract research organization (CRO) hai, kyunki woh ek sath kai sponsors ke documents chalate hain aur isliye logon ko dhang se train karte hain. In-house sponsor roles aam taur par pehle step ke bajaye doosra step hote hain. Freelance regulatory work hota hai lekin shuruat mein mushkil hai, kyunki clients compliance ke baare mein judgement buy kar rahe hote hain aur yahi woh cheez hai jo ek naye writer ke paas abhi nahi hoti.
Jo aap kisi ke aapko hire karne se pehle kar sakte hain, woh hai padhna. ICH E3 aur E6(R3) ko shuru se aakhir tak padhein. Fir E3 section list (jo public hai) aur paper ke data ka use karke ek published trial ka mock synopsis likhein. Yeh single sample kisi bhi certificate se zyada bada demonstration hai, kyunki yeh dikhata hai ki aap ek specification ke andar kaam kar sakte hain. Iske saath hi AMWA guide to regulatory writing career ka ek sahi aur free overview hai.
<ProTip title="🧰 Toolkit:" description="Learn advanced styles, cross referencing, and track changes in your word processor properly before your first role. Regulatory writers spend more time in those features than in any other tool" />
Guideline Hi Job Description Hai
Agar isme se koi ek idea bahar le jaane layak hai, to woh yeh hai ki regulatory writing koi aisi mysterious specialism nahi hai jise aap sirf andar reh kar hi seekh sakte hain. Specifications published, free aur readable hain. Ek competent regulatory writer ko beginner se jo cheez alag karti hai, woh secret templates ka access nahi hai, balki relevant guideline ko haal hi mein itne achhe se padhna hai ki yeh pata ho ki woh sach mein kya kehti hai, na ki woh jo har koi assume karta hai.
<CTA title="Bring Order to Long Technical Drafts" description="Structure, draft, and check complex documents without losing consistency between sections" buttonLabel="Try Jenni Free" link="https://app.jenni.ai/register" />
Ush document type se shuru karein jo aapko sabse pehle milne ki umeed hai, jo aam taur par ek protocol summary, ek consent form, ya ek lay summary hota hai. Iske guideline ko padhein, ek likhein, aur kisi aise vyakti se check karwayein jo ise roz karta hai. Iski underlying discipline, yaani methods ko precisely describe karna, ek consistent formal register banaye rakhna, aur har claim ke peeche source ko point karne ki ability, is field mein har jagah ek jaisi hai.